Post-Brexit, Great Britain and the EU are separate regulatory corridors. Eclevar runs in-vitro diagnostic clinical performance studies as a UK legal entity, under ISO 20916, with HRA and REC submissions via IRAS, MHRA notification and NHS access across all four nations, so one protocol supports both UKCA marking and an IVDR Annex XIII Performance Evaluation Report.
 UK legal entity · In-house CRAs
One accountable UK team runs your IRAS submission, NHS site set-up and performance study, on the MILO Studio platform that won the Platinum Award at the xShare & EUCROF 2026 Open Call.
Horizon Europe · Grant Agreement No. 101136734 · Amsterdam, 2 Feb 2026
Diagnostics and device leaders choose Eclevar for UK and EU performance evidence. Read all client success stories.



An IVD clinical performance study is a prospective investigation that generates the clinical evidence a diagnostic device needs: proof that it measures what it claims, in the intended population, to the required accuracy.
In-vitro diagnostics are regulated differently from devices. In the UK they fall under UK MDR 2002 (as amended), route to market via UKCA marking, and require an MHRA performance study notification. Study conduct follows ISO 20916, and the evidence must support a performance evaluation built on three pillars: scientific validity, analytical performance and clinical performance, captured to the standard an IVD CRO is built to deliver.
Because Great Britain and the EU are now separate corridors, many manufacturers face duplicate studies. Eclevar avoids that with a single-protocol design that yields evidence for both a UKCA submission and an IVDR (EU) 2017/746 Annex XIII Performance Evaluation Plan and Report, exported from one clean database rather than reconstructed after the fact.
Access to patients runs through the NHS: HRA and REC approval via IRAS, NHS Trust Confirmation of Capacity and Capability, and devolved-nation permissions, all coordinated by our feasibility and site selection and clinical data management teams on MILO Studio.
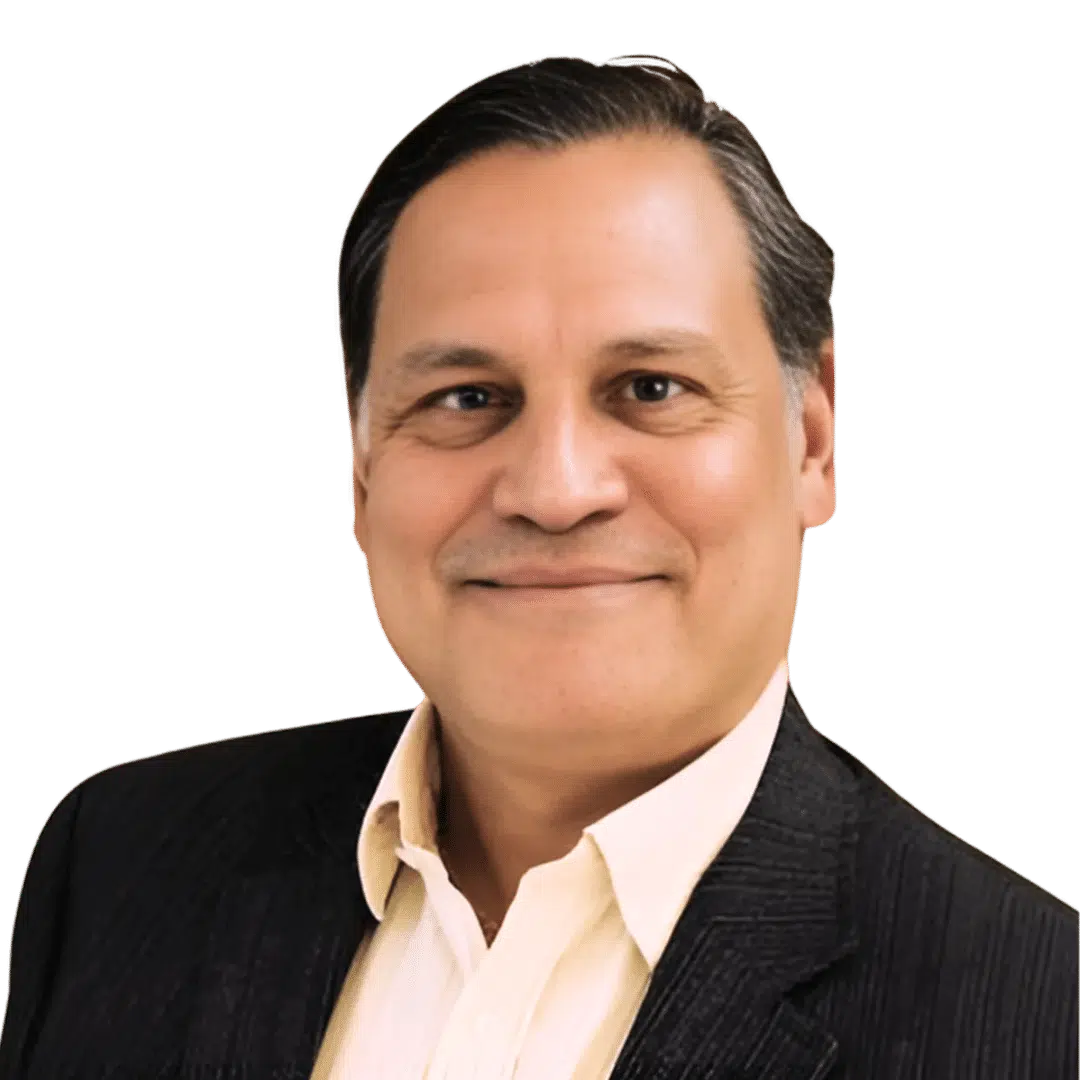

Eclevar's model brings the regulatory perspective inside the CRO, eliminating the gap between what manufacturers submit and what assessors expect. Dr Mark Da Costa, COO and former TÜV SÜD reviewer, and Dr Nikhil Khadabadi, CMO and also a former TÜV SÜD senior reviewer, lead a team that designs IVD performance evidence exactly the way UKCA assessors and Notified Bodies expect to read it, from the first protocol version.
Who runs your IRAS submission, and from where, decides how fast you reach NHS patients and whether one study serves both markets. The gap between a generalist offshore CRO and a UK-resident IVD team is structural.
From performance evaluation strategy to database lock, every activity is run to ISO 20916 and captured with full traceability for UKCA assessment and Notified Body review.
Classification, performance study route and an IVDR-aligned Performance Evaluation Plan, set with our regulatory affairs and clinical strategy team before the first submission.
In-house IRAS applications, HRA and REC liaison, and MHRA performance study notification under UK MDR 2002, run in parallel to protect the timeline.
Site feasibility, contracting and Confirmation of Capacity and Capability across the four UK nations, driven by our feasibility and site selection and study initiation teams.
Risk-based monitoring and in-house clinical data management on an EDC built for diagnostic accuracy endpoints with ALCOA+ traceability.
Single-database exports that support both a UKCA submission and an IVDR (EU) 2017/746 Annex XIII Performance Evaluation Report, with UK data residency.
UK Responsible Person services for non-UK manufacturers, backed by our ISO 13485 quality system for inspection readiness.
One accountable team covers every stage of the performance study lifecycle, so nothing is lost between a protocol, an NHS site and a UKCA or IVDR submission.
Prospective performance studies under ISO 20916, from feasibility and site selection and study initiation through HRA/REC submissions and risk-based monitoring.
Performance Evaluation Plan and Report to IVDR Annex XIII, structured on scientific validity, analytical and clinical performance and reviewed by former Notified Body assessors.
100% in-house clinical data management and EDC: eCRF design, DMP and DVP on MILO Studio, with ALCOA+ scoring and UK data residency.
Regulatory affairs and clinical strategy for UKCA marking and IVDR, with MHRA notification and UK Responsible Person support.
Post-market performance follow-up and real world evidence to keep performance data current after market entry.
Every study run under a quality management system certified to ISO 13485, structured for UKCA assessment and Notified Body review.
An IVD manufacturer needed UK clinical performance data for a UKCA submission but also sold into the EU and faced the prospect of a second, duplicate study for the IVDR. Working as a UK IVD CRO, Eclevar designed a single protocol to ISO 20916 that anticipated both regulations from inception: scientific validity, analytical performance and clinical performance endpoints mapped once, to serve both a UKCA package and an IVDR Annex XIII Performance Evaluation Report.
In-house IRAS submissions and direct HRA liaison moved the study through approval, NHS sites were contracted across more than one UK nation, and data were captured on MILO Studio with ALCOA+ traceability and UK data residency, exported as dual-pathway datasets from one clean database.
Outcome: one study instead of two, a defensible UKCA and IVDR evidence base, and no scramble to retrofit a single-market dataset to the other corridor.
Former Notified Body reviewers from TÜV SÜD. We know what fails IVDR and UKCA assessment and design performance evaluations to avoid it.
In-house IRAS submissions, direct HRA liaison and UK-resident CRAs reaching NHS sites the same week, not offshore handoffs.
Proprietary EDC built for diagnostic accuracy endpoints, with ALCOA+ scoring and dual-pathway exports.
One protocol to ISO 20916 that serves both a UKCA submission and an IVDR Annex XIII Performance Evaluation Report.
Platinum Award winner, xShare & EUCROF 2026, recognized for clinical validation and data interoperability.
ISO 13485 certified. Every performance study structured for UKCA assessment and Notified Body inspection readiness.
A UK legal entity with employed CRAs and project leads, not subcontractors, coordinated with our EU and Japan corridors, gives sponsors one accountable team from IRAS submission to IVDR Performance Evaluation Report.
Whitepapers, client testimonials and publications produced by our teams and partners (BSI, TÜV SÜD, RegenLab).

Written with Notified Body BSI: a practical reading of the clinical evidence expectations under EU MDR 2017/745, with lessons that carry across to IVDR performance evaluation and any high risk device program.
Eclevar manages RegenLab's PMCF program on chronic wound devices. This is a randomized study of 160 patients across 14 sites in 5 EU countries, covering both diabetic foot ulcer (DFU) and venous leg ulcer (VLU) indications. The partnership combines Eclevar's ISO 14155 clinical expertise with the Milo Studio platform to deliver post-market clinical follow-up evidence that supports both Notified Body scrutiny and reimbursement endpoints, from protocol design through to final study report.
« Eclevar, with its tailor-made approach and advanced Milo Studio platform, represents a major strategic asset. »

Most interventional IVD performance studies require MHRA review under UK MDR 2002 (as amended) alongside HRA and REC approval. Studies using genuinely left-over, fully anonymised residual samples can sometimes follow a lighter HRA-only route, but the classification must be confirmed early because it drives the whole submission timeline.
Recent commercial studies have run to a published median of roughly 73 days for combined HRA and HCRW approval, with NHS Trust Confirmation of Capacity and Capability typically adding around 35 days. A clean IRAS application prepared to ISO 20916 is the single biggest lever on that timeline.
Yes, when the protocol is designed for both regulations from the start. A single-protocol architecture can generate scientific validity, analytical performance and clinical performance evidence that supports a UKCA submission and an IVDR (EU) 2017/746 Annex XIII Performance Evaluation Report from one database. Retrofitting a single-market study to the other pathway rarely works cleanly.
Non-UK IVD manufacturers placing devices on the Great Britain market must appoint a UK Responsible Person who liaises with the MHRA and holds documentation on the manufacturer's behalf. Eclevar can act as your UK Responsible Person and run the performance study as one connected service.
The UK is a four-nation research environment. A study is submitted through IRAS and coordinated across NHS Research Scotland, Health and Care Research Wales and the Northern Ireland HSC R&D Division, with devolved-nation permissions managed in parallel so recruitment is not held up in any one territory.
IVD performance studies are governed by ISO 20916 for study conduct, IVDR (EU) 2017/746 Annex XIII for the performance evaluation and UK MDR 2002 (as amended) for the UKCA route, supported by ISO 13485 quality management, UK GDPR and ICH E6(R3) good clinical practice. Performance evaluation is built on scientific validity, analytical performance and clinical performance.
A UK legal entity with in-country employed staff can make IRAS submissions, liaise directly with the HRA and reach NHS sites the same week, rather than working through subcontractors. A specialist IVD CRO with former Notified Body reviewers designs the performance evaluation the way UKCA assessors and Notified Bodies expect to read it.
Send us your protocol synopsis and talk to our UK specialists about a single-protocol study built to ISO 20916 for UKCA and IVDR from day one.
Book a scoping call