
EU MDR · PMS · PMCF
Post-market quality and compliance, done properly.
Under EU MDR, a CE mark is not the finish line. Post-market surveillance and PMCF are what keep a device safe, compliant and on the market. Here is how the system fits together.
European Champion
Platinum Award 2026
Eclevar MedTech & Milo Health · xShare × EUCROF Open Call
Led by authority
Post-market, from people who assess it.
A post-market system is only useful if a Notified Body accepts it. Ours are built by clinicians and former reviewers.
Pierre-Marie Boutanquoi
CERs and PMCF plans under EU MDR and MEDDEV 2.7/1 Rev 4, and Notified Body deficiency response.
in LinkedIn
Dr Mark Da Costa
Cardiac surgeon and former lead Notified Body reviewer at TÜV SÜD. 400+ devices CE-certified.
in LinkedInAwards, funding, accountability
Europe’s best-rated medical device CRO.
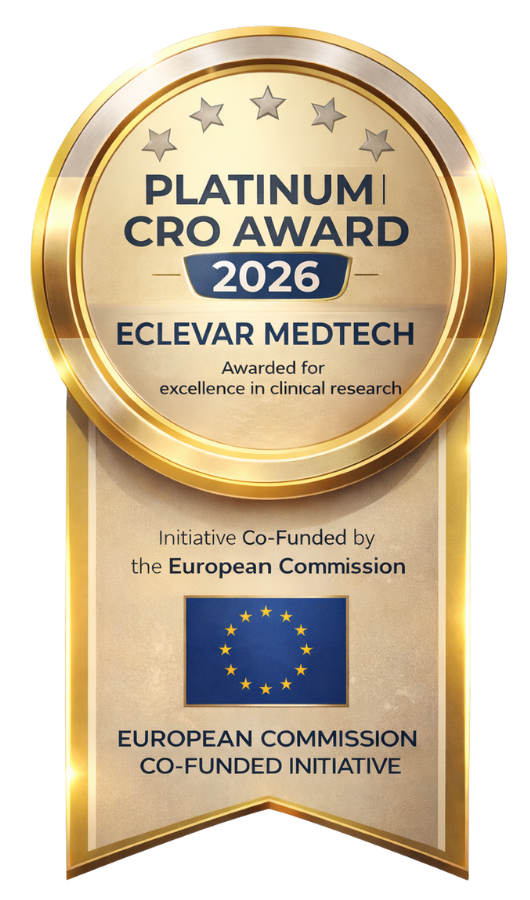
Platinum Award 2026
Top tier at the xShare × EUCROF Open Call, awarded to Eclevar MedTech and its Milo Health platform, presented at EUCROF 2026 in Amsterdam.
The announcement →Co-funded by the European Union
Selected through the xShare Open Call for clinical research innovation, Horizon Europe.
xShare results →Independently reported
Distinction confirmed by an independent third party, the CVBF, also an awardee of the xShare × EUCROF Open Call.
CVBF coverage →The system
PMS and PMCF, and how they connect.
EU MDR treats the post-market phase as a continuous loop: you plan surveillance, gather data proactively, analyze it, and feed the conclusions back into your technical documentation.
01
What post-market surveillance is
Post-Market Surveillance (PMS) is the systematic process of actively collecting and reviewing experience from devices on the market. Its purpose is to detect problems early, confirm the benefit-risk ratio remains acceptable, and drive corrective action when needed, across the whole life of the device.
02
PMS vs PMCF
PMS is the umbrella; PMCF is its proactive, clinical arm.
The distinction
- PMS gathers all post-market experience: complaints, vigilance, literature, sales
- PMCF proactively generates new clinical data on a CE-marked device
- Reactive signals alone are rarely enough for higher-risk devices
- PMCF confirms safety and performance rather than waiting for problems
03
The reporting loop
The data has to go somewhere. Findings feed the Periodic Safety Update Report (PSUR), update the Clinical Evaluation Report (CER) and the risk management file (ISO 14971), and, where relevant, the IFU. A post-market system that collects data but never updates the file is a common, and serious, deficiency.
04
Getting it accepted
The winning approach is a PMS plan whose outputs map directly to the documents a Notified Body reviews, with a PMCF plan sized to the device’s risk class. We design the loop so every post-market signal has a defined route into your technical documentation.
Post-market at a glance
A loop, not a one-off.
Talk to a specialist
Is your post-market system Notified-Body-ready?
Book a free scoping call. We build a PMS and PMCF system whose outputs map to the documents a reviewer will check.
Book a free scoping callA CE mark you don’t maintain is a CE mark at risk.
An expert read builds the PMS and PMCF system that keeps your device compliant, and turns post-market data into evidence a Notified Body accepts.
Book a free scoping callQuestions we hear first